 Europe Regulatory Services
Medical Device Regulatory Consultancy Services for Europe
- EU MDR 2017/745
- EU IVDR 2017/746
- CE marking
Our Services
Our CE Marking Services for the EU
Select a service to see the requirement in detail and how MEDREG supports you.
EUDAMED Registration and Compliance
European Database on Medical Devices (EUDAMED) is an IT system established by Regulation (EU) 2017/745 on medical devices and Regulation (EU)…
 Learn more
Unique Device Identification (UDI) Management
Unique Device Identification (UDI) is a regulatory system established under EU MDR 2017/745 and IVDR 2017/746 to ensure the unique identification and…
 Learn more
Facility, Product & Directive to Regulation Gap Analysis
Gap analysis is a process that looks at where an organization is now and compares it to where it wants to be.
 Learn more
Detailed Risk Management
Risk management is a systematic process used throughout the medical device lifecycle to identify hazards, estimate and evaluate associated risks…
 Learn more
European Authorized Representative Services
An Authorised Representative (AR) is a natural or legal person established within the European Union who is formally appointed by a non-EU…
 Learn more
Person Responsible for Regulatory Compliance (PRRC) Services
The Person Responsible for Regulatory Compliance (PRRC) is a key requirement under Article 15 of the EU Medical Device Regulation (EU MDR 2017/745)…
 Learn more
General Safety and Performance Requirements (GSPR) as per Annex I
The General Safety and Performance Requirements (GSPR), outlined in Annex I of EU MDR 2017/745 and IVDR 2017/746, define the fundamental safety…
 Learn more
Technical Documentation as per Annex II
Before a medical device can be placed on the European Union market, the manufacturer must establish and document its conformity with the applicable…
 Learn more
Post-Market Surveillance (PMS) as per Annex III
Post-Market Surveillance (PMS) is a systematic, ongoing process of collecting, monitoring, and evaluating data on the safety and performance of a…
 Learn more
Detailed Clinical Evaluation and its documentation
As per MEDDEV 2.7/1 Revision 4, Clinical Evaluation is defined as: A systematic and planned process to continuously generate, collect, analyse and…
 Learn more
Post-Market Clinical Follow-up (PMCF)
Post-Market Clinical Follow-Up (PMCF) is a proactive process for collecting and evaluating clinical data on medical devices after they have been…
 Learn more
Summary of Safety and Clinical Performance (SSCP) and Summary of Safety and Performance (SSP)
The Summary of Safety and Clinical Performance (SSCP) is a regulatory requirement under Regulation (EU) 2017/745 (EU MDR) for Class IIb implantable…
 Learn more
Notified body selection, communication and MDR codes
A Notified Body is an independent third-party conformity assessment organization designated by a European Union Member State to evaluate whether…
 Learn more
Risk-Based Device Classification and WET Assessment
Medical device classification under Regulation (EU) 2017/745 (EU MDR) is a systematic, risk-based process that determines the regulatory class of a…
 Learn more
Conformity Assessment Procedures
A conformity assessment is the regulatory process used to demonstrate that a medical device or IVD complies with the applicable requirements of the…
 Learn more
Quality Management System (QMS) Requirements
A Quality Management System (QMS) is fundamental requirement for medical device manufacturers seeking access to the European market.
 Learn more
Cleaning and Disinfection validation
Reusable medical devices play a critical role in modern healthcare by supporting multiple surgical procedures throughout their service life.
 Learn more
Sterilization Validation and Bioburden Testing
Sterilization validation is a systematic and documented process used to demonstrate that a sterilization method can consistently achieve the required…
 Learn more
Biocompatibility evaluation
Biological evaluation is a systematic, risk-based process used to determine the potential for an unacceptable adverse biological response resulting…
 Learn more
Usability engineering evaluation
Usability engineering is an essential aspect of medical device safety, focusing on the interaction between the device, its intended users, and the…
 Learn more
Technical Documentation Grouping
MEDREG provide expert services for technical documentation grouping under EU Regulations, helping manufacturers systematically categorise medical…
 Learn more
Implant Card
An implant card is a patient-specific identification document intended to provide patients with essential information about an implanted medical…
 Learn more
Vigilance Data Collection and Monitoring
Medical device vigilance is a regulatory process for identifying, reporting, investigating, and managing serious incidents and field safety…
 Learn more
Systems and Procedure Packs
Under Regulation (EU) 2017/745 (EU MDR), systems and procedure packs are combinations of medical devices and, where permitted, other products…
 Learn more
Steps to Get a CE Mark
To obtain CE marking, manufacturers must follow an appropriate conformity assessment route based on the classification and risk level of their medical device. The selected route determines the extent of Notified Body involvement and the regulatory requirements that must be met. Obtaining CE marking requires manufacturers to follow a structured conformity assessment process that demonstrates compliance with applicable European regulations. The process typically involves the following steps:
Define the Intended Purpose
Clearly establish the device's intended use, target population and clinical application.
Identify Applicable Regulatory Requirements
Determine whether the device is governed by MDR or IVDR and identify relevant harmonized standards, common specifications, and MDCG guidance documents.
Qualification of device as a Medical Device
As per MDR / IVDR Article 2.
Classify the Device
Assign the appropriate risk class according to the classification rules in the applicable regulation.
Select the Conformity Assessment Route
Choose the conformity assessment procedure based on the device classification and regulatory requirements.
Implement a Quality Management System (QMS)
Establish and maintain a compliant QMS covering design, manufacturing, and post-market activities.
Develop the Device and Technical Documentation
Design, manufacture, validate, and document the device to demonstrate compliance with regulatory requirements.
Select an Appropriate Notified Body
Identify a Notified Body designated under the applicable MDR or IVDR scope and competent to assess the device technology, classification, and conformity assessment route. Early engagement supports efficient certification planning and assessment timelines.
Submit a Pre-Application to the Notified Body (NB)
Provide preliminary information for verification of device scope, classification, and preparation of a conformity assessment quotation.
Lodge the Formal Application
Submit the required QMS and technical documentation, together with the formal application and supporting information.
Application Review and Contract Agreement
The NB reviews the application for completeness and eligibility, and upon acceptance, establishes a written agreement with the manufacturer.
Conformity Assessment
The NB performs the required audits, technical documentation is reviewed thoroughly i.e. GSPR as per Annex I, Technical documentation as per Annex II, Post-Market Surveillance information as per Annex III, and any applicable consultation procedures.
Annex IX - Quality Management System (QMS) and Technical Documentation
Most commonly used conformity assessment route and is intended for manufacturers that have implemented a comprehensive Quality Management System (QMS). It involves an assessment of the QMS and a review of the device's technical documentation by a Notified Body. Depending on the device class, the level of technical documentation review may vary.
Annex X - Type Examination
Focuses on the evaluation of a representative device sample. The Notified Body examines and tests the sample, along with the associated technical documentation, to verify compliance with MDR requirements, particularly the General Safety and Performance Requirements (GSPRs).
Annex XI - Product Conformity Verification
Annex XI provides additional routes for demonstrating product conformity. It may be used in combination with Annex X or, for certain lower-risk devices, alongside technical documentation. This route includes assessment of production quality assurance systems and/or verification of specific product batches to confirm compliance.
Regardless of the conformity assessment route selected, manufacturers must need to prepare and maintain technical documentation in accordance with Annex II and establish Post-Market Surveillance (PMS) documentation as required by Annex III. These documents serve as the foundation for demonstrating ongoing compliance throughout the device lifecycle.
Final Review and Certification Decision
The NB independently reviews the assessment results, verifies resolution of non-conformities, and decides whether certification can be granted.
Issue the EU Declaration of Conformity and Affix the CE Mark
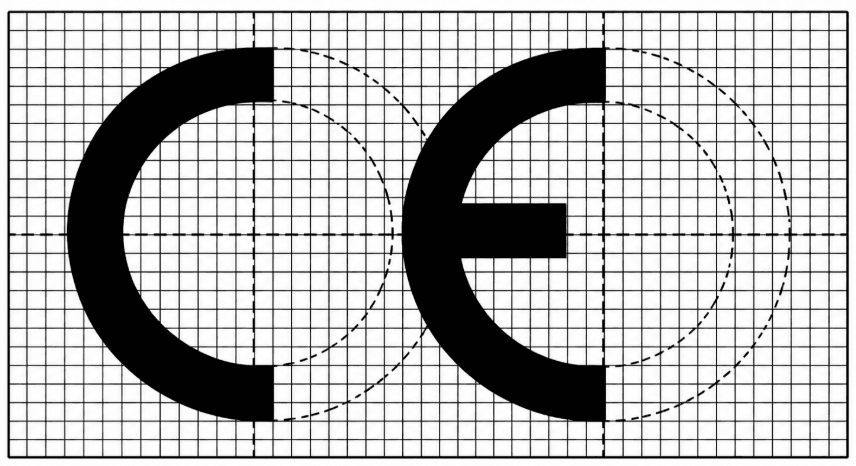
Following successful certification, the manufacturer prepares the EU Declaration of Conformity and affixes the CE marking in accordance with Annexes IV and V of the MDR/IVDR. Where applicable, the Notified Body identification number shall accompany the CE mark. The CE marking must comply with the prescribed proportions and have a minimum height of 5 mm, unless waived for small-scale devices, demonstrating conformity with regulatory requirements and enabling placement of the device on the European market
Certificate Issuance and Post-Certification Surveillance
NB issues the MDR/IVDR certificate and conducts ongoing surveillance activities to verify continued compliance throughout the certificate validity period.
MEDREG helps manufacturers to understand and meet the applicable requirements of the EU Medical Device Regulation (EU MDR 2017/745) and the In Vitro Diagnostic Medical Devices Regulation (EU IVDR 2017/746).
From device classification and technical documentation to conformity assessment and Notified Body coordination, we provide structured guidance throughout your CE marking journey.
Our goal is to help you build a strong regulatory foundation for safe, compliant and successful entry into the European market. In accordance with EU MDR 2017/745 and EU IVDR 2017/746, we offer complete regulatory consulting services to assist medical device manufacturers in obtaining and retaining the CE mark.
Our CE marking services are designed for
- Medical device manufacturers seeking European market access.
- IVD manufacturers preparing for EU IVDR compliance.
- Medical device manufacturers preparing for EU MDR compliance.
- Manufacturers transitioning from legacy European directives to the applicable MDR or IVDR requirements.
- Manufacturers preparing for Notified Body assessment.
- Manufacturers requiring technical documentation gap assessments.
- Manufacturers seeking support with clinical evaluation, performance evaluation, or biological evaluation.
- Manufacturers looking to strengthen their QMS and post-market compliance processes.
MEDREG - Your Trusted Partner for CE Marking & European Market Access
Why MedReg
Why choose MedReg
Expert consultants, thorough regulatory insight, and a seamless path to compliance, that’s our promise for your medical devices.
Project Strategy & Gap Assessment
Before a single page is drafted, we map your device to the right regulations and find every gap in your files, so your documentation is built once, built right and submission-ready.
Simplifying Complexity
We decode Complex regulatory requirements and define a clear, actionable pathway, to ensure your projects move forward efficiently and compliantly.
Comprehensive Understanding
We dive deep into your requirements, identifying all critical factors to ensure our solutions address every aspect of your regulatory needs.
Documentation You Can Trust
As experts, we provide clear, accurate, and accessible documentation, keeping you informed at every stage of your project.
Consultation
Compliance Made Simple – Start with a Free Consultation
Confidential & Audit-Proof
Request a consultation
Direct confidential advisory with senior CDSCO, CE MDR, and FDA specialists.